Why Choose Testsealabs Rapid Test for Global Buyers?
Global buyers need more than a fast result. They need confidence in product quality, documentation, supply continuity, and responsible use. Testsealabs Rapid Test is designed for professional testing environments where clear procedures and dependable performance matter. Its value should be assessed through evidence, not attractive packaging or ambitious promises.
A practical evaluation begins with the details. Buyers should review intended use, sample requirements, detection windows, storage conditions, lot identification, and available technical documents. Clear instructions can reduce handling mistakes in busy clinics, laboratories, and distribution centers. Consistent packaging also helps teams recognize expiration dates and batch information quickly. These small details matter.
Quality, however, is never a single feature. Test performance may vary with sample collection, timing, user training, and the population tested. No rapid test is perfect. That matters. Buyers should examine validation data, quality-control procedures, certifications, and local regulatory requirements before placing an order. Independent professional judgment remains essential, especially when results are unclear or unexpected.
For international customers, reliable communication is equally important. Responsive support can clarify shipping conditions, storage temperatures, documentation, and complaint procedures. A capable supplier should explain limitations openly, not hide them behind marketing language. Testsealabs Rapid Test may support efficient screening workflows, but buyers must match each product with an appropriate application. The strongest purchasing decision combines technical evidence, responsible distribution, and realistic expectations. That approach protects users, strengthens trust, and supports dependable healthcare practices across different markets.
What Is Testsealabs Rapid Test and How Does It Work?
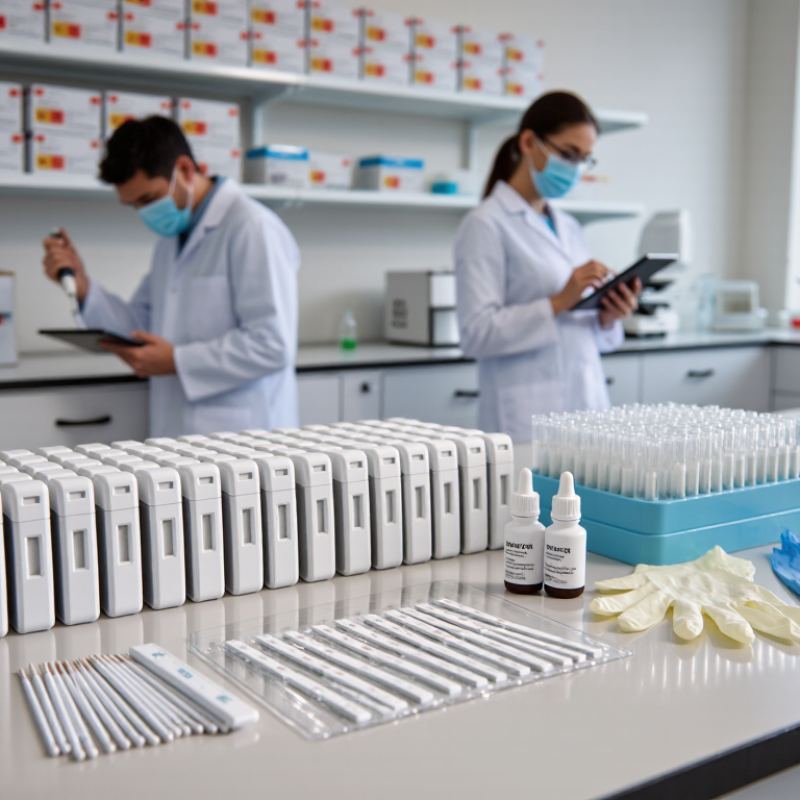
A rapid diagnostic test is a portable screening tool built for speed and simple handling. It usually uses a lateral-flow strip inside a sealed cassette. A user adds a prepared sample, often nasal fluid, to the test well. The liquid moves across the membrane through capillary action. If target antigens are present, labeled antibodies form a visible line. A separate control line confirms that the liquid flowed correctly.
The World Health Organization’s target product profile for antigen tests expects at least 80% sensitivity and 97% specificity. These figures explain why test performance matters for global buyers. Results often appear within 15 to 30 minutes, but timing varies by product and sample quality. In practical use, clear instructions, stable storage, and trained handling reduce avoidable errors. Yet rapid does not mean perfect. Testing too early, collecting too little sample, or reading outside the stated time can produce misleading results. I have seen simple workflow mistakes affect confidence more than the device itself.
Tips: Check the expiration date, storage temperature, and control line. Use a timer. Do not interpret a faint or late line casually. WHO and FIND reports also emphasize independent validation before procurement. A rapid test can support fast decisions, but unexpected results may require laboratory confirmation. That extra step feels inconvenient, but skipping it creates a blind spot.
Why Choose Testsealabs Rapid Test for Global Buyers? - What Is Testsealabs Rapid Test and How Does It Work?
| Data Dimension | Typical Rapid-Test Information | Relevance for Global Buyers |
|---|---|---|
| Test format | Most rapid tests use lateral-flow technology. The device uses a sample pad, reagent area, membrane, test line, control line, and absorbent pad. | The format supports visual result reading and is suitable for decentralized testing without complex laboratory equipment. |
| Primary analyte | Depending on the assay, the test may detect pathogen antigens, antibodies produced by the body, or nucleic acids. | Buyers should match the analyte to the intended purpose, testing stage, and applicable local regulations. |
| Common specimen types | Possible specimens include nasal or throat swabs, saliva, urine, or capillary whole blood, depending on the specific assay. | Specimen requirements affect user training, collection materials, logistics, and market registration. |
| Typical time to result | Many lateral-flow assays provide a visual result within approximately 10–30 minutes. The exact reading time is defined in the instructions for use. | Short processing time can support screening in clinics, workplaces, pharmacies, schools, travel settings, and remote areas. |
| How the reaction works | The specimen is applied to the test strip. If the target is present, it binds with labeled reagents and forms a visible line at the test region. The control line confirms that the test has flowed correctly. | A clear, easy-to-follow mechanism helps reduce operational complexity and supports use by trained non-laboratory personnel. |
| Result interpretation | A valid test normally requires a visible control line. A test line may indicate a detected target, while no control line means the result is invalid and the test should be repeated. | Packaging should include clear diagrams, reading times, invalid-result instructions, and language appropriate for the target market. |
| Analytical sensitivity | Rapid antigen tests generally detect higher target concentrations than molecular tests. Performance can vary with specimen quality, timing, and target concentration. | Buyers should review product-specific sensitivity data, study design, population, and intended-use claims rather than relying on a generic percentage. |
| Specificity considerations | Specificity describes how well a test identifies samples without the target. Cross-reactivity and interfering substances can affect results. | Independent validation data and interference studies are important for assessing reliability in the intended market. |
| Storage and transport | Lateral-flow products are commonly supplied in sealed foil pouches and usually require controlled temperature and humidity conditions specified by the manufacturer. | Temperature limits, shelf life, shipping conditions, and stability data should be checked before international procurement. |
| Required equipment | Visual lateral-flow assays generally require the test cassette, collection device, buffer when applicable, timer, and appropriate personal protective equipment. | Minimal equipment can reduce infrastructure requirements and simplify deployment in resource-limited settings. |
| Quality documentation | Important documents may include instructions for use, performance evaluation, batch records, certificates of analysis, stability data, and quality-management documentation. | Complete documentation supports distributor due diligence, customs clearance, tender submissions, and regulatory review. |
| Regulatory suitability | Authorization requirements differ by country and by intended use. A product may require local registration, authorized representation, labeling compliance, and post-market monitoring. | Global buyers should confirm the applicable regulatory pathway and verify that claims, packaging, and documentation match the destination market. |